The silver bullet
Aging processes can reinforce one another. A precise intervention at a shared causal link might let several systems recover. Finding such a link is the silver-bullet hypothesis.
Explore the selection
The search for a large effect from one intervention
A silver bullet starts with a hypothesis about ageing itself: changing one pivotal process might restore several body systems at once. The proposals below explore different ways this could happen. Each must test whether a local intervention can produce a wider, lasting benefit.
A decisive experiment must separate the proposed mechanism from competing hypotheses. An improvement shared by all predictions tells us less than a result that reveals which mechanism produced it. The collection therefore considers both the promised reach of an intervention and the explanatory value of its proposed test.
Published experiments provide the starting points. Their species, tissues and measured outcomes matter: a result in a mouse organ supports a narrower claim than rejuvenation of a whole human body. Under each proposal, the literature explains what has been observed and which step remains a hypothesis.
Three ways to change the course
Stop the source
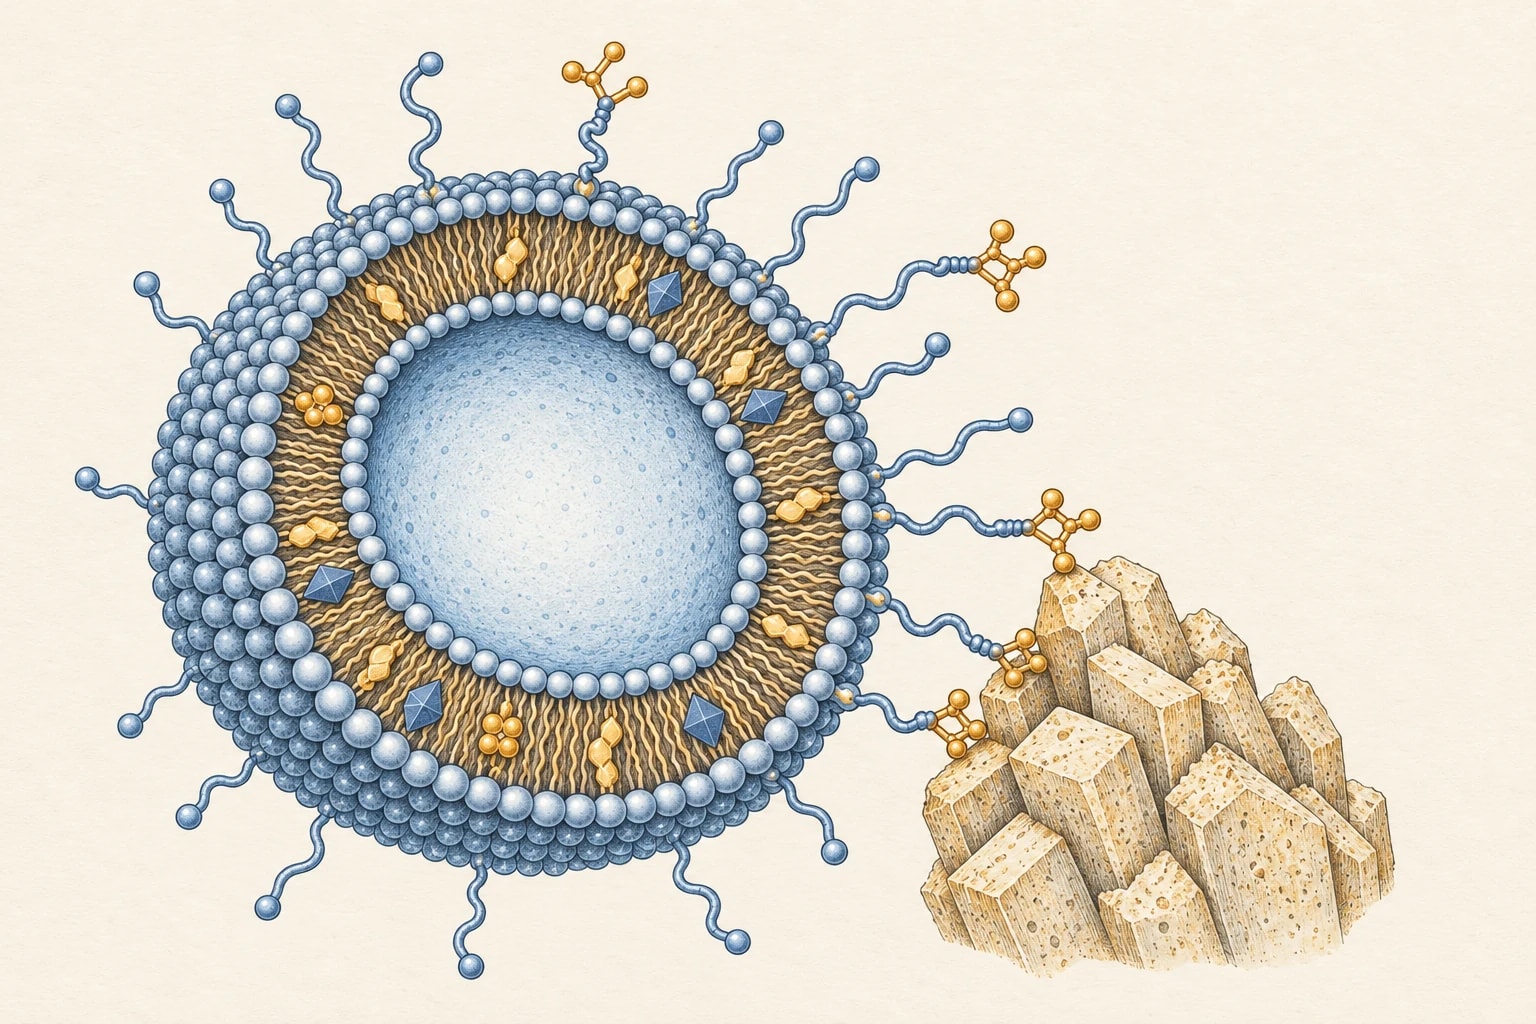
Interrupt a signal that keeps causing damage. If tissue still has repair capacity, stopping the source may let it recover.
Alendronate on the liposome surface binds bone mineral. The carrier delivers dasatinib and quercetin; this formulation was studied in mouse models of treatment-induced bone loss.
doi:10.1016/j.fmre.2024.12.010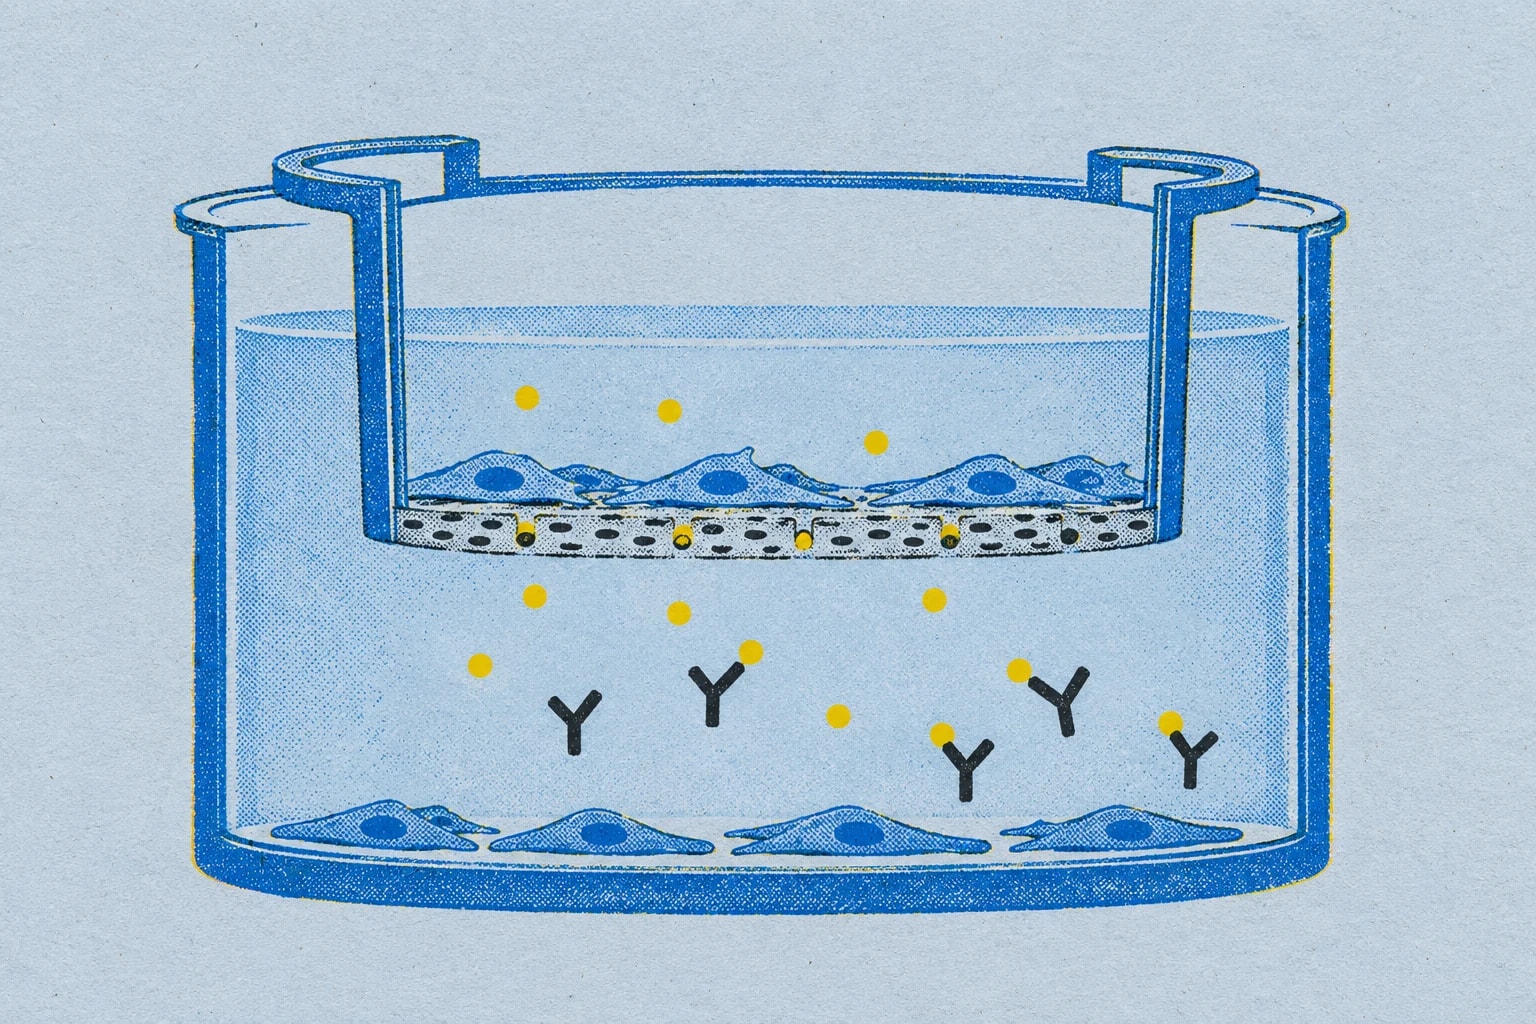
Break the cycle
Break a feedback loop in which damage and its consequences keep sustaining each other. A brief intervention could then have a lasting effect.
Proposed test of soluble signals: a porous membrane separates source and recipient cells, while antibodies neutralize a selected signal in the medium. Persistence of the altered state must be measured separately.
Release repair
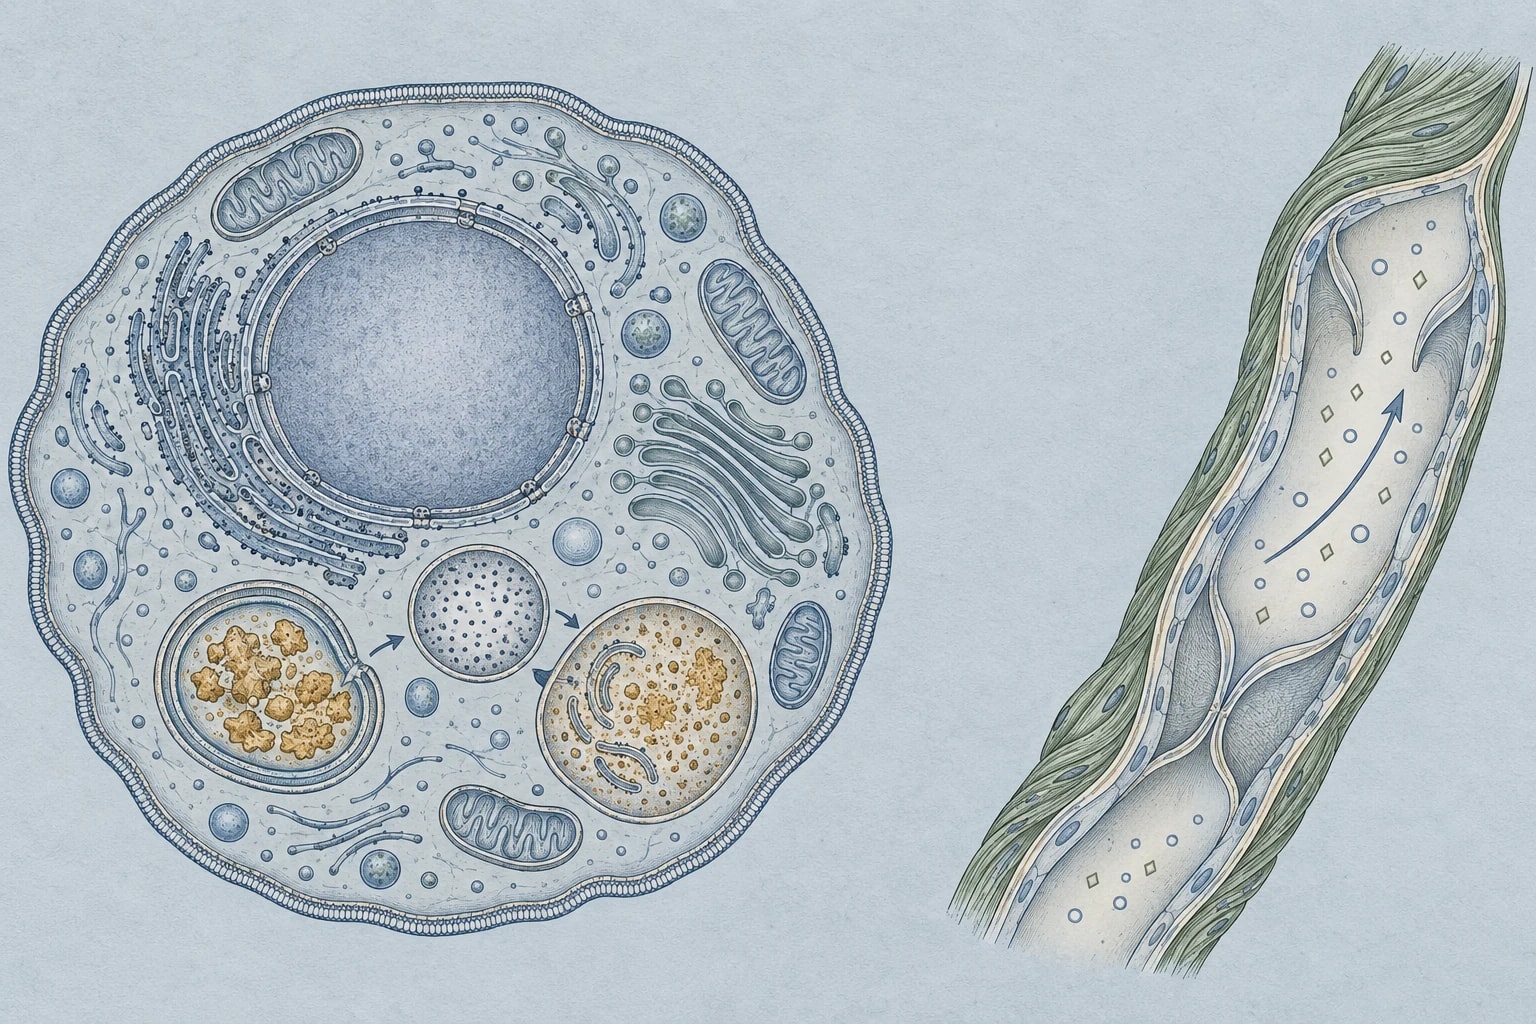
Remove the bottleneck holding back existing repair, such as a limited route for removing waste from tissue.
Two processes for an independent experimental comparison: intracellular degradation through autophagosomes and lysosomes, and transport through a collecting lymphatic vessel. Their proposed interaction remains to be tested.
These are research strategies. Their effects depend on the particular mechanism and tissue.
Hypotheses worth testing
This collection brings together published experimental starting points and hypotheses proposed by our system. The system-generated selection changes as assessments and experimental designs change. Deferred candidates remain in the record with the reason for setting them aside. Inclusion means the proposal merits a discriminating test; it does not establish efficacy.
Hypothesis 1 · Proposed by our system
Preserve muscle and brain function through localized liver treatment
What this could change
A treatment confined to a small liver cell population could preserve strength, memory and thinking during aging. If restoring ammonia capture in these cells also extends remaining life, treatment development could focus on one target for several age-related impairments.
- Glutamine synthetase as a central element in hepatic glutamine and ammonia metabolism: novel aspects.Full text available
- Glucagon Receptor Inhibition Reduces Hyperammonemia and Lethality in Male Mice with Urea Cycle Disorder.Full text available
- Taurine transporter (TauT) deficiency impairs ammonia detoxification in mouse liver.Full-text access unverified
- Pharmacokinetic and Pharmacodynamic Properties of L-Ornithine L-Aspartate (LOLA) in Hepatic Encephalopathy.Full text available
Proposed hypothesis
Restoring glutamine synthetase activity in 80% of mapped liver cells near veins could reduce ammonia peaks and preserve muscle and brain function. Normalized ammonia with unchanged functional decline and survival would reject the proposed route to longer life.
What the experiment would decide
The decisive pattern would be lower ammonia peaks after meals and activity, preserved strength, memory and thinking, and longer remaining life after selective restoration in 80% of the target cells. Recreating the original ammonia pattern should remove the functional benefits. If the processes targeted by competing interventions remain unchanged, this would favor ammonia mediation over attributing these benefits to changes in those processes. Other interventions could still provide independent benefits. This result would support developing localized liver treatment to preserve functions across organs.
The target sustains the mutual amplification of dysfunctions
The hypothesis and the observation that tests itWhat the studies establish
Glutamine synthetase incorporates ammonia into the amino acid glutamine. A 2021 review, accessible only as an abstract, identifies a small population of liver cells around central veins as essential for maintaining low ammonia levels. A 2019 study in mice lacking the taurine transporter supports this pathway while revealing a limitation. At 12 months, an inactivating chemical modification reduced glutamine synthetase activity by 40%, and perfused livers had reduced glutamine-producing capacity. At 3 months, ammonia was already elevated despite normal enzyme abundance and activity; RhBG, a protein involved in ammonia transport into cells, was reduced by about 50%. Urea production remained unaffected in the perfusion experiments. These findings suggest that ammonia entry into cells can limit capture even when the enzyme works normally. The accessible study text has limited coverage and describes no selective restoration experiment.
In a 2021 study, an antibody blocking the glucagon receptor reduced elevated ammonia and increased survival in male Otcspf-ash mice with an inherited urea-cycle defect aggravated by a high-protein diet. Treatment increased liver glutamine synthetase expression, reduced expression of glutaminase, which breaks down glutamine, and increased circulating glutamine, lean mass and body mass. This links altered nitrogen metabolism with survival under disease stress, but does not establish that ammonia reduction caused the survival benefit or that correction confined to the proposed cells would suffice. Lean mass also differs from muscle strength. Only the abstract is accessible; it gives no treatment duration, follow-up period or numerical effect sizes.
A 2019 review of L-ornithine L-aspartate describes relevant muscle and brain outcomes in liver disease. In patients with cirrhosis, elevated ammonia and muscle wasting, a single infusion did not restore muscle metabolic responsiveness, whereas infusions over 7 days restored the response to a protein meal and inhibited protein breakdown. In rats with acute liver failure caused by interruption of hepatic blood supply, treatment normalized plasma ammonia, delayed severe neurological complications and reduced brain edema. The treatment acts through several pathways, including urea production and glutamine synthesis outside the liver, so these findings do not isolate the proposed cell population. Separately, experiments in mouse, pig and human intestinal tissue found that ammonia increased nerve-driven contractions and reduced relaxations; inhibiting glutamine synthetase potentiated ammonia's effects in mouse experiments. That study's limited accessible text concerns intestinal responses and cannot establish preservation of skeletal muscle strength, memory or thinking.
The required fraction of corrected cells needs its own evidence. An abstract describing explanted livers from two women with ornithine transcarbamylase deficiency reports that approximately 20-30% of hepatocytes expressing that functional enzyme was insufficient for metabolic stability. This concerns a different enzyme and cell population, so the percentage cannot establish a glutamine synthetase restoration threshold. Mouse lineage tracing found that labeled zone 3 hepatocytes largely retained their zonal distribution over 50 weeks under normal conditions, while chronic injury could change their distribution and metabolic identity. This was a broader population than the proposed glutamine synthetase-positive subset. The hypothesis predicts insufficient benefit after restoration in 40% of the initially mapped target cells, sufficient benefit at 80%, and no additional benefit at 100%. These are untested predictions. Establishing equivalence between 80% and 100% requires a predefined criterion; these three levels cannot locate an exact minimum between 40% and 80%.
What remains to be established
Selective restoration has not been shown to preserve function or extend life during ordinary aging. Elevated ammonia despite normal enzyme activity in young taurine-transporter-deficient mice indicates that enzyme restoration alone may be insufficient. Survival evidence concerns male mice with inherited disease; human treatment evidence concerns cirrhosis, and acute brain protection in rats does not demonstrate rejuvenation. A 2026 cirrhosis study predicted metabolic effects in liver, brain and muscle through computational modeling; it did not test recovery after treatment. A nephrology review accessible only as an abstract describes ammonia handling across several organs and its relationship to acid-base balance. Interpreting ammonia reversal requires separating accompanying glutamine and acid-base changes. Verified selective restoration that normalizes ammonia while leaving function and survival unchanged would reject the claimed sufficiency under the tested conditions. Benefits persisting after a valid reversal would challenge ammonia mediation. The proposed experiment leaves species, starting age and observation period unspecified. Durability, long-term safety and relevance to human aging remain unestablished.
- Glucagon Receptor Inhibition Reduces Hyperammonemia and Lethality in Male Mice with Urea Cycle Disorder.Full text available
- Taurine transporter (TauT) deficiency impairs ammonia detoxification in mouse liver.Full-text access unverified
- Integrative host-microbiome modeling uncovers the implication of oral-gut translocation in advanced cirrhosis.Full text available
- Pharmacokinetic and Pharmacodynamic Properties of L-Ornithine L-Aspartate (LOLA) in Hepatic Encephalopathy.Full text available
- Hyperammonemia Physiology and Management: A Nephrology-Centered Review.Abstract only
Hypothesis 2 · Proposed by our system
Lasting protection for vessels, muscles and liver
What this could change
A short treatment could protect blood vessels, muscles and liver long after it ends if closing persistent breaks in genetic material removes a continuing source of inflammation. Confirmation would support pursuing longer remaining life through repair that introduces some new sequence errors. Those errors would be an acceptable cost only if lasting benefits outweigh their harmful consequences.
Proposed hypothesis
In old organisms, brief polymerase theta activity may seal persistent DNA (deoxyribonucleic acid) breaks, reducing inflammatory secretion and extending remaining life despite added mutations. Lasting benefit without new DNA junctions would refute the mechanism.
What the experiment would decide
The decisive pattern in old mice would be persistent new DNA joins with additional insertions and deletions, fewer physical breaks and weaker damage transmission between organs at one, three and six months after treatment activity has demonstrably ended. These changes should accompany better vascular, muscle and liver function and ultimately longer remaining life. The proposed comparison predicts no benefit from catalytically inactive polymerase and less durable benefit from brief inflammatory suppression despite similar early improvement. This pattern would favor break closure over simple inflammatory suppression and strengthen it relative to hypotheses involving tissue rearrangement, infection or damaged proteins. Researchers could then prioritize closing persistent breaks for lasting organ protection. Lasting benefit without new joins would contradict the proposed mechanism; failure of functional recovery or life extension would count against the corresponding prediction only after adequate closure of the relevant targeted breaks had been established.
The target sustains the mutual amplification of dysfunctions
The hypothesis and the observation that tests itWhat the studies establish
Polymerase theta, encoded by POLQ, helps join broken ends of deoxyribonucleic acid (DNA), the molecule carrying genetic information. Reviews published in 2016 and 2022 describe joining through short matching sequences, which can introduce insertions or deletions. This repair can prevent larger deletions in some settings, but its outcome depends on the structure and processing of the broken ends. Increasing the enzyme alone therefore cannot be assumed to close persistent breaks. Related repair machinery also supports cancer cells: in cultured human U2OS cancer cells depleted of the repair protein BRCA2, reducing XRCC1, a protein that organizes repair complexes, impaired the restart of DNA copying and increased sensitivity to replication stress while reducing chromosome abnormalities. These experiments demonstrate a tradeoff between cell survival and genetic instability in that setting; they did not test polymerase-theta enhancement in aging.
- XRCC1 promotes replication restart, nascent fork degradation and mutagenic DNA repair in BRCA2-deficient cells.Full text available
- DNA polymerase θ (POLQ), double-strand break repair, and cancer.Full-text access unverified
- To indel or not to indel: Factors influencing mutagenesis during chromosomal break end joining.Full-text access unverified
A 2020 EMBO Reports study describes immediate cessation of cell proliferation after persistent double-strand breaks, followed days later by the inflammatory secretory program associated with cellular senescence. It also reports that histone deacetylase inhibitors induced this program without DNA damage. The response required noncanonical functions of damage-response proteins on chromatin, the complex of DNA and associated proteins. This challenges the assumption that inflammatory secretion necessarily requires continuing breaks: secretion might persist after their closure. Access is limited to the abstract and synopsis, which do not specify the cellular model or provide detailed methods and results. They establish damage-independent secretion, but not what happens after physical break closure.
- Non-canonical ATM/MRN activities temporally define the senescence secretory program.Full-text access unverified
Welch and colleagues tested inflammatory communication from damaged mouse neurons. Medium collected 24 hours after etoposide treatment and washout altered the morphology of microglia, the brain's resident immune cells, during a six-hour exposure of brain slices. Removing CCL2 or CXCL10, signaling proteins released by the neurons, alleviated several of these changes. In CK-p25 mice, with neurodegeneration induced at three to four months of age, suppressing neuronal p65, a regulator of inflammatory genes, reduced microglial proliferation at one, two and six weeks and preserved synaptic density at six weeks. Neuronal loss, amyloid accumulation and tau phosphorylation were unchanged; the early comparison also found unchanged numbers of neurons positive for a DNA-damage marker. These findings support harmful signaling from damaged neurons and show that partial tissue protection can follow signaling suppression without demonstrated physical repair.
A 2025 study found persistent intestinal changes in male mice exposed at eight to ten weeks of age to 50 centigray of simulated galactic cosmic radiation. Five months later, intestinal DNA-damage and senescence markers were elevated, inflammatory cytokine expression had increased, and nutrient-transporter expression and blood biomarkers of intestinal injury were altered. The study measured activation of a DNA-sensing inflammatory pathway but did not experimentally manipulate that pathway or test break repair. It documents coexistence of lasting damage and inflammation after irradiation. Reversal through break closure and damage transmission between organs remain hypotheses.
What remains to be established
Whether briefly increasing polymerase theta can close naturally persistent breaks and extend remaining life in old organisms remains unestablished. A mouse study found unchanged lifespan after loss of polymerase nu, encoded by POLN, a different enzyme. Its separate POLQ comparison found that radiation-associated damage-marker foci in spermatids, developing sperm cells, declined to similar levels by five hours. These marker measurements do not establish physical closure of persistent breaks. A 2026 polymerase-theta review, accessible only as an abstract, describes both genome protection and cancer-associated rearrangements; its abstract reports no aging-treatment result. Apparent repair must be distinguished from loss of damaged cells. Harmful mutations, chromosome-end fusions and tumors could outweigh early benefits; breaks at telomeres, the protective chromosome ends, require separate assessment. The proposed experiment lacks a quantitative criterion for adequate closure. Disease-model observations over weeks and radiation follow-up over five months establish neither systemic rejuvenation nor human benefit. The proposed mathematical transition from amplification to damping of damage remains unmeasured, and local damping alone would not establish longer life.
- DNA polymerase θ (POLQ), double-strand break repair, and cancer.Full-text access unverified
- Neurons burdened by DNA double-strand breaks incite microglia activation through antiviral-like signaling in neurodegeneration.Full text available
- Analysis of DNA polymerase ν function in meiotic recombination, immunoglobulin class-switching, and DNA damage tolerance.Full text available
- cGAS/STING Pathway Mediates Accelerated Intestinal Cell Senescence and SASP After GCR Exposure in Mice.Full text available
- Polymerase theta: Genome protection through regulated deployment.Abstract only
Hypothesis 3 · Proposed by our system
One repair target for lasting heart and muscle recovery
What this could change
If mitochondrial renewal alone can sustain recovery of the heart and skeletal muscles and extend life, treatment development could concentrate on restoring this process in surviving cells. Establishing that senescent-cell removal and suppression of inflammatory secretion deliver their lasting benefits through the same process would help determine when direct renewal could replace those interventions.
Proposed hypothesis
Mitochondrial renewal in surviving cells may explain all lasting benefits of the tested interventions. Benefit from cell clearance despite selectively blocked renewal, or failure of renewal alone with senescent cells and their inflammatory secretion held near baseline, would reject the claim.
What the experiment would decide
The hypothesis predicts that selectively suppressing mitophagy in surviving heart and skeletal-muscle cells eliminates the lasting functional and lifespan benefits of senescent-cell removal and reduced inflammatory secretion, although those interventions still achieve their immediate targets. Restoring mitophagy should restore the benefits. Direct mitochondrial renewal must also produce those benefits while senescent-cell numbers and secretion remain near their starting levels. Together, these outcomes would favor one sufficient repair route over competing hypotheses of independent benefits or an obligatory combination. Researchers could then prioritize direct renewal under the tested conditions. Retained benefits during a valid selective blockade, or failure of verified renewal alone in a valid sufficiency test, would contradict the central claim and redirect research toward other targets or combinations.
The target limits the body's capacity to repair
The hypothesis and the observation that tests itWhat the studies establish
In a 2016 study, oral spermidine started at 18 months increased median mouse lifespan by approximately 10%. Separate experiments in aged mice found reduced cardiac hypertrophy, improved filling during relaxation and increased mitochondrial respiration. Spermidine also enhanced autophagy, intracellular breakdown and recycling, including mitophagy, the selective disposal of mitochondria. In young mice lacking Atg5, a protein required for autophagy, specifically in heart muscle cells, spermidine failed to protect the heart and aggravated hypertrophy. This supports an autophagy requirement for cardioprotection under those conditions. It does not isolate mitophagy or establish what caused lifespan extension. The authors explicitly considered additional contributions to longevity; that uncertainty alone does not refute the hypothesis.
- Cardioprotection and lifespan extension by the natural polyamine spermidine.Full text available
A 2026 skin study provides mechanistic support and a challenge to universal dependence on mitophagy. Substances secreted by human umbilical-cord mesenchymal stem cells improved skin barrier measurements, collagen preservation and cellular-senescence markers in five-month-old female mice during 40 days of ultraviolet exposure, with treatment during the final 30 days. Ultraviolet-exposed immortalized human keratinocytes also showed protection after 48 hours of treatment. Mdivi-1, used to suppress mitophagy, weakened protection. Adding H151, which blocks the inflammatory signaling protein STING, restored reductions in inflammatory cytokine levels and restored the senescence-associated protein lamin B1 while mitophagy-related measurements remained suppressed, in both cells and mice. This pharmacological rescue supports protection through downstream inflammatory control despite impaired mitochondrial recycling. It challenges universal necessity, but does not test lasting heart or muscle recovery or lifespan.
Muscle studies connect functional improvement with several simultaneous changes. In male SAMP10 accelerated-aging mice, an infusion of human umbilical-cord mesenchymal stromal cells improved muscle performance at the reported three-month assessment, alongside better mitochondrial structure, increased markers of mitochondrial formation, and reduced inflammation and programmed cell death. The article contains inconsistent descriptions of follow-up timing. A study of the herbal preparation Juyuanjian reports improved muscle mass and strength in SAMP8 mice and reduced muscle-fiber damage in nematodes. Experiments also reported enhanced mitochondrial formation, reduced inflammation and suppression of mechanisms promoting muscle-protein breakdown. Only the abstract is available, restricting assessment of methods, causal tests and follow-up. A 2026 review describes human urolithin A trials reporting altered mitochondrial gene expression in muscle after four weeks and improved muscle endurance during four-month treatment. These findings motivate testing how much recovery mitochondrial renewal mediates. Its sufficiency for all benefits remains unestablished; the review's proposed additional benefits from combining interventions also remain hypothetical.
- Natural Bioactive Compounds Targeting Key Hallmarks of Aging: Functional Food Potential of Spermidine, Fisetin, Berberine, and Urolithin A.Full text available
- Juyuanjian attenuates sarcopenia through dual regulation of the Akt/FoxO1 and SIRT1/PGC-1α pathways.Abstract only
- Human umbilical cord-derived mesenchymal stromal cells ameliorate aging-associated skeletal muscle atrophy and dysfunction by modulating apoptosis and mitochondrial damage in SAMP10 mice.Full text available
Research in other tissues supports communication between cells as a way to influence mitochondrial maintenance. Small membrane vesicles released by mechanically stimulated human mesenchymal stem cells improved survival and mitochondrial-quality-control measurements in oxidatively stressed rat cartilage cells. Inhibiting miR-330-3p, a regulatory molecule carried by the vesicles, weakened these effects. Weekly joint injections improved repair of surgically created cartilage defects in rats assessed at four and eight weeks. These experiments support a contribution from the vesicle cargo, but do not establish mitochondrial renewal alone as sufficient for repair. A 2026 review of aging mesenchymal stem and stromal cells describes functional improvements after mitochondrial interventions and emphasizes that delivery of mitochondrial material to lysosomes, cellular degradation compartments, does not prove completed breakdown. Neither work establishes that engulfing removed senescent cells triggers renewal in neighboring heart or skeletal-muscle cells.
What remains to be established
Mitochondrial renewal remains unestablished as both necessary and sufficient for these lasting benefits. Senescent cells persist after stable arrest of division; the proposed connection between their immune removal, engulfment of cells or their remains, and renewal in neighboring cells requires a separate causal test. Mitophagy provides disposal, whereas renewal also requires replacement. Blocking recycling can erase treatment benefits by injuring surviving cells, and experimentally maintaining cell numbers or secretion can itself distort the sufficiency test. Both confounders require exclusion. The skin rescue is the closest experimental challenge, but concerns different tissues, endpoints and timescales. Cellular findings, rodent experiments and short human trials do not establish systemic rejuvenation or human lifespan extension. The decisive experiment still lacks a specified species, follow-up duration and quantitative success criterion.
- The Secretome Derived From Human Umbilical Cord Mesenchymal Stem Cells Improves Skin Photoaging by Enhancing Mitophagy to Inhibit the cGAS-STING Pathway.Full text available
- Natural Bioactive Compounds Targeting Key Hallmarks of Aging: Functional Food Potential of Spermidine, Fisetin, Berberine, and Urolithin A.Full text available
- Mitochondria-Lysosome Quality Flux in Aging Mesenchymal Stem/Stromal Cells: Quality-Control Bottlenecks and Therapeutic Opportunities for Regenerative Decline.Full text available
Hypothesis 4 · Proposed by our system
Reduce lung and kidney damage while preserving infection defense
What this could change
If this hypothesis holds, treatment could reduce chronic inflammatory damage to the lungs and kidneys while preserving the enhanced infection defense established by earlier immune exposures. Targeting a harmful signal from a small population of blood-forming stem cells could allow useful immune memory in their descendants to remain intact.
- Mesenchymal Stromal Cells Facilitate Neutrophil-Trained Immunity by Reprogramming Hematopoietic Stem Cells.Full text available
- BCG HSP70 Reprograms Macrophages via Central Trained Immunity to Suppress Prostate Cancer.Full text available
- Rare epigenetic alterations are conserved across hematopoietic differentiation stages after mycobacterial infection.Full text available
Proposed hypothesis
Aging blood-forming stem cells may sustain damage by secreting interleukin-6 while their descendants retain antimicrobial memory. Insufficient secretion, or damage declining only after descendant replacement following selective shutdown in stem cells, would reject the mechanism.
What the experiment would decide
The decisive result would be lower interleukin-6 between infections and less lung and kidney damage after selectively stopping its production in stem cells once immune memory has formed. Improvement should precede replacement of existing descendants, with their early infection control preserved. Restoring interleukin-6 delivery at the measured physiological rate should restore damage. This pattern would favor a continuing damaging signal and weaken a mechanism requiring descendant replacement. Research could then focus on suppressing stem-cell secretion while preserving protective memory. Insufficient stem-cell output, benefit only after descendant replacement, or impaired infection control would redirect research toward other sources or linked programs of damage and defense.
The target sustains the mutual amplification of dysfunctions
The hypothesis and the observation that tests itWhat the studies establish
Trained immunity is a lasting change in how innate immune cells respond to subsequent challenges. A 2025 review describes mouse transplantation and human vaccination studies linking its persistence to changes in blood-forming stem and progenitor cells. The authors also propose that hyperresponsive descendants could sustain inflammation in postinfectious illness, while acknowledging that this mechanism remains unestablished in myalgic encephalomyelitis/chronic fatigue syndrome. This identifies a competing hypothesis for persistent inflammation: cytokine release by descendants.
In a Mycobacterium avium study, mice receiving blood-forming stem and progenitor cells from previously infected donors were challenged three months after transplantation. Four weeks later, they had fewer bacteria in the spleen than controls; no mortality difference was detected. Only a few infection-associated changes in chromatin, the packaging that regulates access to genes, were shared across developmental stages. After transplantation and 12 weeks away from the infected environment, infection-associated enrichment of H3K4me3, a chemical mark on chromatin proteins, no longer persisted despite retained functional immune memory. Separately, macrophages derived from trained progenitors released more interleukin-6, an immune signaling protein, after bacterial challenge. These findings support durable protection, challenge permanent storage of memory in that particular mark, and demonstrate cytokine production by descendants.
Another mouse study used Dnak, a heat-shock protein from a mycobacterial vaccine strain. Seven days after donor treatment, a population containing blood-forming stem cells and early progenitors was transferred into lethally irradiated recipients, which were challenged with Staphylococcus aureus seven days later. Cells from donors given 1 or 2 mg of Dnak reduced splenic bacterial counts at 16 hours and improved survival during 25 days of follow-up; 0.2 mg produced no significant improvement in these endpoints. This demonstrates transferable protection against acute infection. In separate experiments, macrophages derived from treated mice released more interleukin-6 after exposure to a bacterial component or tumor cells. Protective training can therefore accompany increased inflammatory signaling by descendants.
A 2023 transplantation study exposed mouse blood-forming stem and progenitor cells to substances released by human stromal cells, which support other cells in tissues. Some stromal cells had first been stimulated with a synthetic mimic of bacterial genetic material. Months after transplantation, recipients of cells trained with this preparation had fewer lung bacteria 24 hours after Pseudomonas aeruginosa infection than control recipients; the reduction in splenic bacterial burden was not statistically significant. Baseline circulating cytokines were comparable between groups at 12 weeks, and interleukin-6 was lower in the trained group after infection. Enhanced defense thus coexisted with comparable baseline cytokines and lower postinfection interleukin-6. The study did not selectively suppress stem-cell interleukin-6 or establish its reduction as the cause of improved bacterial clearance.
What remains to be established
Whether rare, long-lived stem cells directly release enough interleukin-6 between infections to sustain aging-related lung and kidney damage remains unestablished. The closest counterevidence is protection with comparable baseline cytokines or lower postinfection interleukin-6, alongside increased release by trained macrophages in other models. Early improvement and restoration of damage after interleukin-6 replacement cannot alone exclude rapid mediation through existing descendants or the marrow environment. Direct measurements of total stem-cell output and verified intervention specificity are essential. Preserved early defense would support only short-term separation; protection through repeated infections and descendant renewal requires testing. The principal experiments concern mice over days to months. Human vaccination observations and cell-line experiments do not establish this intervention in older people. The proposed experiment leaves species and duration unspecified; systemic rejuvenation and lifespan extension remain unproven.
- Advocating the role of trained immunity in the pathogenesis of ME/CFS: a mini review.Full text available
- Mesenchymal Stromal Cells Facilitate Neutrophil-Trained Immunity by Reprogramming Hematopoietic Stem Cells.Full text available
- BCG HSP70 Reprograms Macrophages via Central Trained Immunity to Suppress Prostate Cancer.Full text available
- Rare epigenetic alterations are conserved across hematopoietic differentiation stages after mycobacterial infection.Full text available
Hypothesis 5 · Proposed by our system
Lasting recovery in the gut and blood vessels after a brief intervention
What this could change
A brief intervention in one tissue could allow both the gut and blood vessels to recover durably if repair-driven cell division sustains damage passing between them. Confirmation would focus treatment development on temporarily interrupting that division, with benefits continuing after normal tissue renewal resumes.
Proposed hypothesis
In linked gut and vascular tissue, repair-driven division may cause DNA replication stress and repeated cell death. Damage that persists during confirmed division arrest but stops when lipid oxidation or mineral particle formation is suppressed would reject this mechanism.
What the experiment would decide
The decisive observation would be lasting cessation of recurring damage in both parts of a linked gut–vascular tissue model after a brief pause in division of the receiving tissue’s support cells, with the initiating injury already removed. Recovery should persist after division and tissue exchange resume under the same daily burden; seven daily cycles are proposed as a preliminary criterion. Persistence with unchanged cell proportions would favor division-driven damage over the rival requiring a change in cell composition. Continued spread during confirmed division arrest, followed by cessation when lipid oxidation or mineral-particle formation is suppressed, would favor the corresponding rival and redirect the treatment target. Durable recovery after the pause would prioritize temporary division control for tests in aged animals. Damage returning simply when division resumes would reject the prediction of lasting recovery.
The target sustains the mutual amplification of dysfunctions
The hypothesis and the observation that tests itWhat the studies establish
In mice exposed to repeated low-dose gamma irradiation, deleting Irf5 suppressed thymic lymphoma development. Researchers attributed this partly to reduced death of thymocytes and blood-forming stem cells, which reduced replacement division and genetic damage associated with difficulties copying genetic material. A separate study reported that Puma-deficient mice resisted radiation-induced lymphoma: blood-forming stem and progenitor cells protected from death showed less replacement division and associated damage. These findings support the possibility that the response to cell loss can generate further injury. Both interventions reduced cell death; neither tested a temporary pause in division. The Irf5 study is available only as an abstract. The accessible Puma report contains summary findings, with detailed methods, results and follow-up unavailable.
Deleting Ddx3x specifically in mouse hepatocytes, the liver’s principal working cells, produced cell death and replacement division in female mutants at six weeks of age, alongside genetic damage and signs of difficulty copying genetic material. The study also linked the deletion to reduced levels of repair factors and eventual liver tumors. This provides another setting in which cell loss, division and damage occur together. However, impaired repair offers a direct route to genetic damage, so their coexistence does not establish that replacement division initiated it. Only the abstract is available; it describes no test of damage transmission between tissues or recovery after temporarily stopping division.
A study of cultured human blood-forming stem and progenitor cells found lower activity of two major pathways repairing double-strand breaks in deoxyribonucleic acid (DNA) than in mature peripheral-blood lymphocytes. Diminished signaling through NF-κB, a regulator of gene expression, limited repair-gene expression and was implicated in these differences. This abstract-only evidence identifies repair capacity as another possible contributor to damage in dividing cells. It does not demonstrate that replacement division creates a self-sustaining damage cycle.
Prostaglandin E2, the proposed division-stimulating signal in the hypothesis, suppressed division in experiments with human cord-blood stem and progenitor cells. It reduced cell death caused by taxol or etoposide at 48 hours; separate assays documented reduced division. Exposure to 50 micromolar prostaglandin E2 for longer than two days became toxic in cultures followed for up to eight days. In immunodeficient mice carrying human blood-forming cells, daily administration of its longer-acting analogue did not improve human blood-cell recovery after irradiation at approximately one week. With etoposide, the human-cell proportion increased, but absolute human-cell numbers did not. Combining the analogue with irradiation or etoposide caused severe, nearly fatal diarrhea. These findings challenge any assumption that prostaglandin E2 consistently promotes division; its proposed role in gut and vascular support cells was not tested.
What remains to be established
The studies concern radiation injury, genetically altered mouse liver and human blood-forming cells in culture or transplanted into mice. They establish neither a self-sustaining aging cycle between the gut and blood vessels nor systemic rejuvenation. Release of prostaglandin E2 by dying cells in this proposed cycle, the substances carrying damage between tissues and a threshold for self-sustaining damage remain unestablished. The prostaglandin findings are relevant counterevidence to uniform stimulation of division, but do not directly test the stromal hypothesis. Stopping division could impair necessary repair; apparent improvement could also reflect cell loss, disrupted exchange or effects on competing mechanisms. The named outcome SPV_1 lacks a biological definition. Seven daily cycles would provide preliminary tissue-model evidence only; durable organ benefits, lifespan effects and human efficacy require separate studies.
- NF-κB-dependent DNA damage-signaling differentially regulates DNA double-strand break repair mechanisms in immature and mature human hematopoietic cells.Abstract only
- In vitro and in vivo evaluation of possible pro-survival activities of PGE2, EGF, TPO and FLT3L on human hematopoiesis.Full text available
- DNA Damage, Liver Injury, and Tumorigenesis: Consequences of DDX3X Loss.Abstract only
- Deletion of Irf5 protects hematopoietic stem cells from DNA damage-induced apoptosis and suppresses γ-irradiation-induced thymic lymphomagenesis.Abstract only
- Apoptosis of leukocytes triggered by acute DNA damage promotes lymphoma formation.Full-text access unverified
Hypothesis 6 · Proposed by our system
Preserve neutrophils’ protective activity to help organs recover
What this could change
If delaying neutrophil removal prolongs their protection of tissue, preserving these immune cells during recovery could help muscle, heart and lungs recover together. Confirmation would help determine when retaining the cells is useful and whether administering the vesicles they release could substitute for keeping the cells.
Proposed hypothesis
After suppression of prostaglandin E2 receptor type 2 (EP2), delaying neutrophil uptake may improve muscle, heart and lung recovery by preserving protective vesicles. Failure of neutrophil-specific CD55 loss to abolish the added benefit would reject the proposed dependence on this protein.
What the experiment would decide
With EP2 suppression verified and initial damage comparable, the proposed test asks whether selectively delaying whole-neutrophil engulfment improves muscle, heart and lung recovery beyond EP2 suppression alone. The decisive pattern would be loss of this additional benefit after neutrophil-specific CD55 deletion and its return after purified CD55-positive vesicle delivery. Accelerating cell removal before vesicle formation should worsen recovery. If tied to CD55-dependent complement inhibition, this pattern would favor the protective-vesicle hypothesis over competing hypotheses based on fewer harmful vesicles or greater tissue tolerance, while allowing mechanisms to coexist. Researchers could then prioritize preserving neutrophils’ release of protective vesicles during recovery. No additional recovery, or benefit persisting after verified loss of the CD55-dependent complement effect, would argue against prioritizing delayed cell removal.
The target limits the body's capacity to repair
The hypothesis and the observation that tests itWhat the studies establish
The 2025 Cell study found that human and mouse neutrophils release large extracellular vesicles, membrane-enclosed particles, as the cells age over hours in culture. A protective subset carries CD55, a protein that restrains complement, a system of immune proteins. In female mice aged 8-10 weeks with Staphylococcus aureus pneumonia, purified vesicles delivered into the trachea one hour after infection reduced inflammation and lung injury at 24 hours after infection without impairing bacterial clearance. Neutrophil-specific CD55 deletion increased and prolonged complement activation and worsened lung injury. Transferring vesicles from normal neutrophils into these mice reduced inflammation and lung injury; CD55-deficient vesicles failed to rescue those endpoints. Separate experiments showed that blocking vesicle CD55 reversed suppression of complement activation. Blocking surface signals that protect vesicles from engulfment shortened their persistence and weakened their anti-inflammatory effect. These findings support protection through persistent vesicles; benefit from delaying removal of their producer cells remains a separate hypothesis.
Prostaglandin E2 receptor type 2 (EP2) responds to the chemical signal prostaglandin E2. In the 2012 study, female mice aged 6-8 weeks genetically lacking EP2 had 59.3% mortality over ten days after pneumococcal infection, compared with 92% in controls. At 24 hours, their lungs contained fewer bacteria and more neutrophils. Their isolated alveolar macrophages, immune cells that engulf material in lung airspaces, showed greater bacterial uptake and killing. Improved antimicrobial defense therefore provides a competing hypothesis for EP2-related benefit. The study did not test delayed neutrophil engulfment or CD55-dependent vesicle protection.
Cell experiments show that prostaglandin signaling regulates uptake of other targets. In human macrophage-like THP-1 cells, prostaglandin E2 reduced uptake of heat-killed Clostridium sordellii during a three-hour assay, with evidence favoring prostaglandin E2 receptor type 4 (EP4) over EP2. In the 2022 study, prostaglandin E2 reduced Escherichia coli uptake by human HL-60 cells and freshly isolated human neutrophils; EP2 antagonism partially reversed inhibition in HL-60 cells. The 2015 antagonist study reported dose-dependent enhancement of macrophage-mediated amyloid-beta plaque clearance from brain slices outside the animal. Only its abstract is available, limiting assessment of that result: the slice species and assay duration are unspecified. These studies support context-dependent regulation of uptake; they do not establish whole-neutrophil removal or protective vesicle production.
- EP4 and EP2 receptor activation of protein kinase A by prostaglandin E2 impairs macrophage phagocytosis of Clostridium sordellii.Full text available
- PGE2 inhibits neutrophil phagocytosis through the EP2R-cAMP-PTEN pathway.Full text available
- A selective prostaglandin E2 receptor subtype 2 (EP2) antagonist increases the macrophage-mediated clearance of amyloid-beta plaques.Abstract only
The 2025 human observational study compared 40 adults aged at least 65 with 40 younger adults. Several circulating CD55-positive vesicle populations were lower in the older group. However, neutrophil-derived CD55-positive vesicles showed no statistically significant group difference or correlation with age or blood glucose. This finding does not support an age-related decline in that measured population. The assay used a size gate defined by 0.22-0.88 micrometer beads, whereas the Cell study described vesicles averaging more than 1 micrometer, limiting direct comparability. The human study measured associations; it did not test protective function, complement inhibition or treatment benefit.
What remains to be established
The proposed connection between EP2 suppression, the environment around macrophages and protective vesicle production remains unestablished. Retention of whole cells must be distinguished from changes in vesicle production or clearance and from redistribution of neutrophils between organs. In the Cell study, a combined treatment prolonging neutrophil survival increased vesicle yield, whereas caspase-3 inhibition prolonged survival without increasing production. Longer survival therefore accompanied greater vesicle output only under some interventions. Mice completely lacking complement component C3 died rapidly from bacterial overload during live infection, showing that complement also supplies essential antimicrobial protection. Protective vesicle treatment was studied in acute pneumonia in young mice, with key vesicle kinetics followed through 72 hours. Aging of individual neutrophils over hours differs from organismal aging. The proposed test specifies neither species nor follow-up duration. Sustained recovery across organs, substantial reduction of age-related dysfunction and longer life remain untested.
Hypothesis 7 · Functional outcomes in aged mice · 2026
Immune signals and blood-cell production
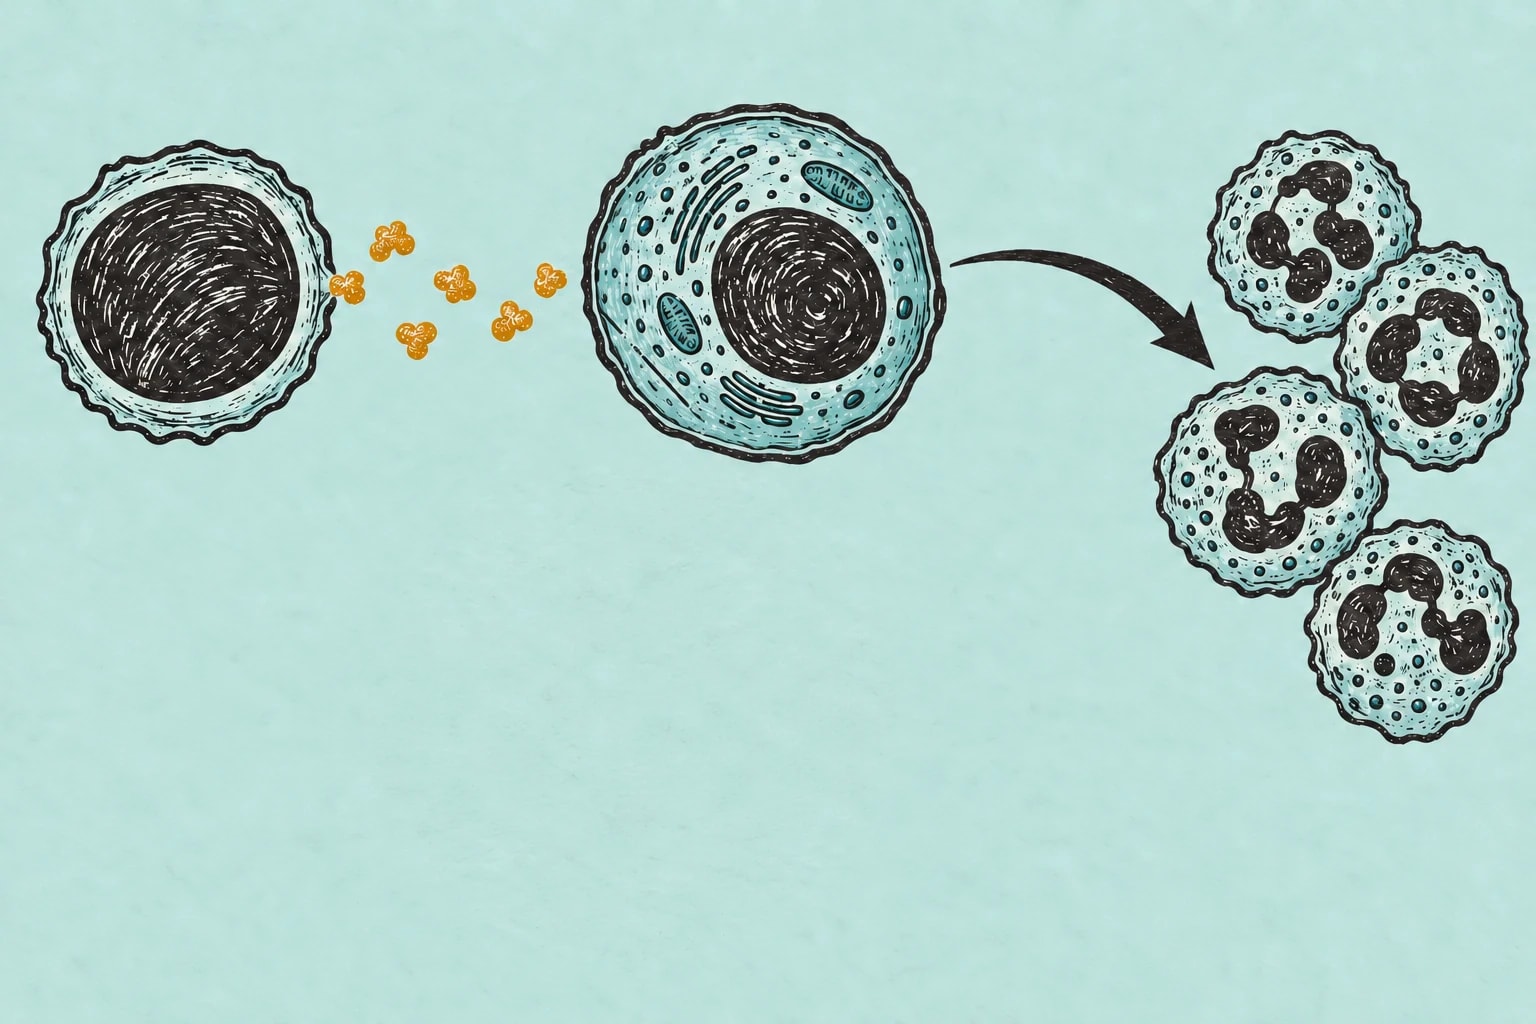
What this could change
Restore blood formation using the body’s own cells
Old blood-forming cells may retain useful capacity that harmful signals prevent them from using. Removing such a signal could make recovery possible while preserving those cells. The case is strengthened by the aged-mouse experiments: blocking the receptor improved strength and coordination as well as blood formation.
What is known
In old mice, mature T cells influenced blood-cell production through CCL5 and CCR5. Blocking CCR5 also improved strength and coordination.
What to test
Do the same old stem-cell clones change how they work when this signal is removed? Benefits to human healthspan remain unproven.
What the studies establish
In aged mice, CD4 T cells in bone marrow released the signaling protein CCL5. Its receptor, CCR5, helped blood-forming progenitors shift toward producing myeloid cells. Deleting the receptor in progenitors weakened this effect, providing a direct test of the proposed signaling pathway.
- Gabandé-Rodríguez et al., Nature Aging, 2026Full text available
In a separate experiment in the same paper, 23-month-old mice received the CCR5 blocker maraviroc for one month. Blood-cell production shifted, and the mice improved in grip-strength and motor-coordination tests. The study therefore connects an identifiable molecular target with physical performance, which makes it a candidate for a broader intervention.
- Gabandé-Rodríguez et al., Nature Aging, 2026Full text available
What remains to be established
These experiments do not establish longer lifespan or lasting recovery after withdrawal. Following individual stem-cell clones would show whether the existing cells change their behavior or the composition of the cell population changes.
Hypothesis 8 · Lifespan effects in worms and flies · 2023
The speed of reading genes
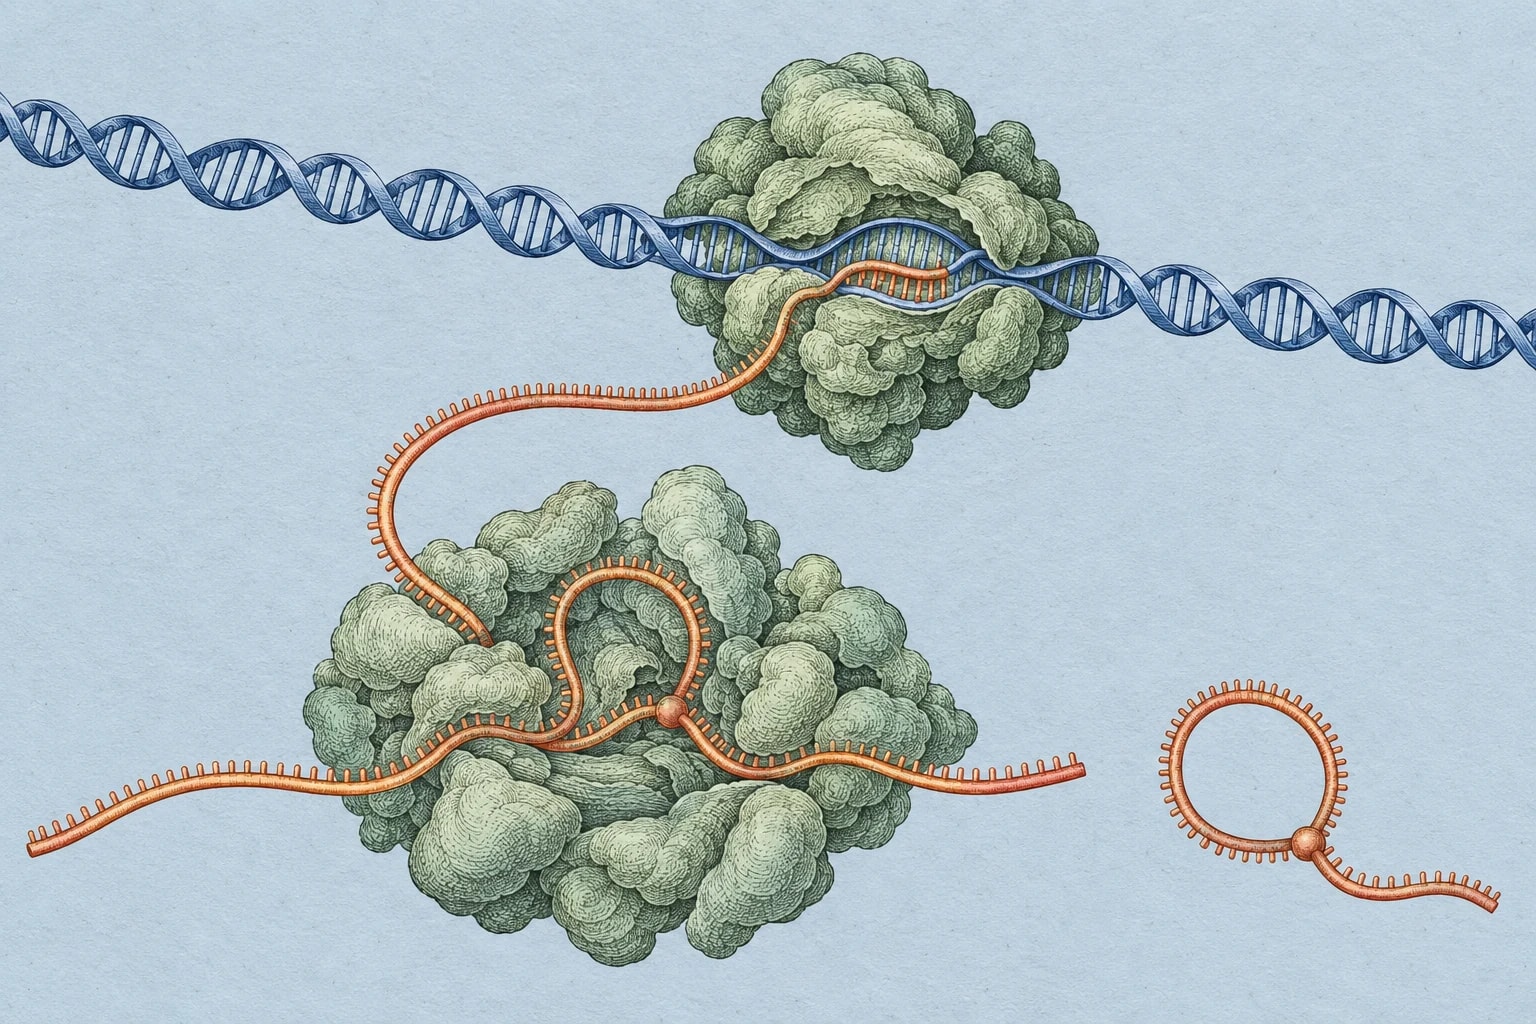
What this could change
One target for changes affecting many genes
Transcription speed affects RNA production and processing across many genes. Correcting it could therefore improve several cellular processes at once. Lifespan extension in worms and flies gives this possibility an experimental basis. Finding a safe intervention that also benefits mammals would establish a shared target for a broad range of age-related changes.
What is known
RNA polymerase II read genes faster with age in samples from five species. Mutations slowing it extended lifespan in worms and flies.
What to test
Separate the effect of elongation speed from a reduction in total RNA production. Benefits in naturally aged mammals require a further experiment.
What the studies establish
Debès and colleagues found that RNA polymerase II, the enzyme that copies DNA into RNA, moved faster on average with age in samples from worms, flies, mice, rats and humans. They also examined changes in RNA processing. This is evidence for a shared age-related process across species; transcription speed did not change in the same direction for every gene.
- Debès et al., Nature, 2023Full text available
Mutations that slowed this enzyme increased median lifespan by about 20% in worms and 10% in flies. Reversing the mutation in worms returned lifespan close to the control level. These interventions make transcription speed a causal candidate, beyond its association with age.
- Debès et al., Nature, 2023Full text available
What remains to be established
The lifespan intervention was tested in worms and flies. A treatment started in naturally aged mammals still needs to show functional benefit while preserving essential RNA production.
Hypothesis 9 · Cell turnover measured in mice · 2019
Removing senescent cells
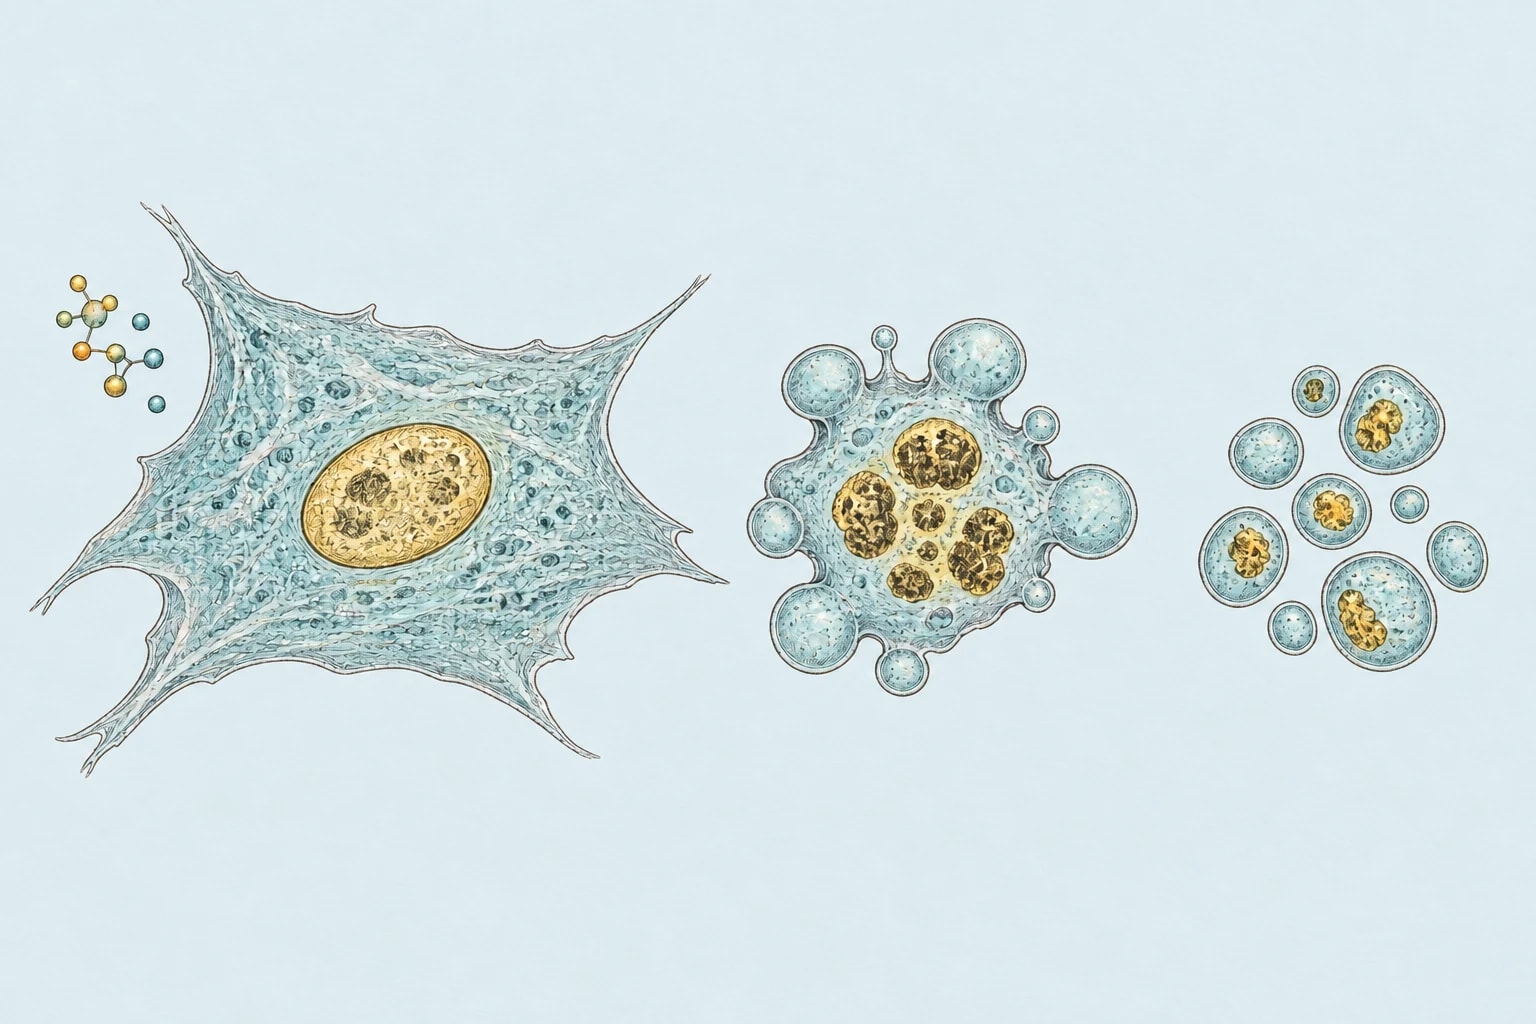
What this could change
Make the effect of cell removal last longer
The model predicts that removing some senescent cells could help the body dispose of the remaining ones more quickly. Treatment could then leave the body better able to limit their accumulation. Confirming this effect would help determine how long the benefit lasts and when another course is needed.
What is known
In old mice, senescent cells accumulated and were removed more slowly. The model links these processes through feedback. Senescent cells have persistently stopped dividing.
What to test
Measure removal and transmission of a persistent dysfunctional state separately. Self-sustaining spread to neighbours remains a distinct hypothesis.
What the studies establish
Karin and colleagues induced senescence in mouse lung cells with bleomycin and followed their disappearance. The estimated half-life was 4.7 days in young mice and 18 days in old mice. Comparing mathematical models with longitudinal measurements supported a feedback in which accumulated senescent cells slow their own removal.
- Karin et al., Nature Communications, 2019Full text available
Ovadya and colleagues tested another part of this mechanism: mice lacking perforin, a protein used by immune cells to kill target cells, accumulated more senescent cells and developed greater tissue damage. Pharmacological removal of senescent cells alleviated some changes. This supports the importance of immune clearance, while recovery of clearance capacity after treatment remains a separate prediction.
- Ovadya et al., Nature Communications, 2018Full text available
What remains to be established
Measurements in injured lungs showed slower removal of senescent cells. The prediction that a short treatment restores the body’s ability to remove these cells requires direct testing after treatment ends.
Hypothesis 10 · Fluid drainage restored in aged mice · 2024
The drainage route outside the skull
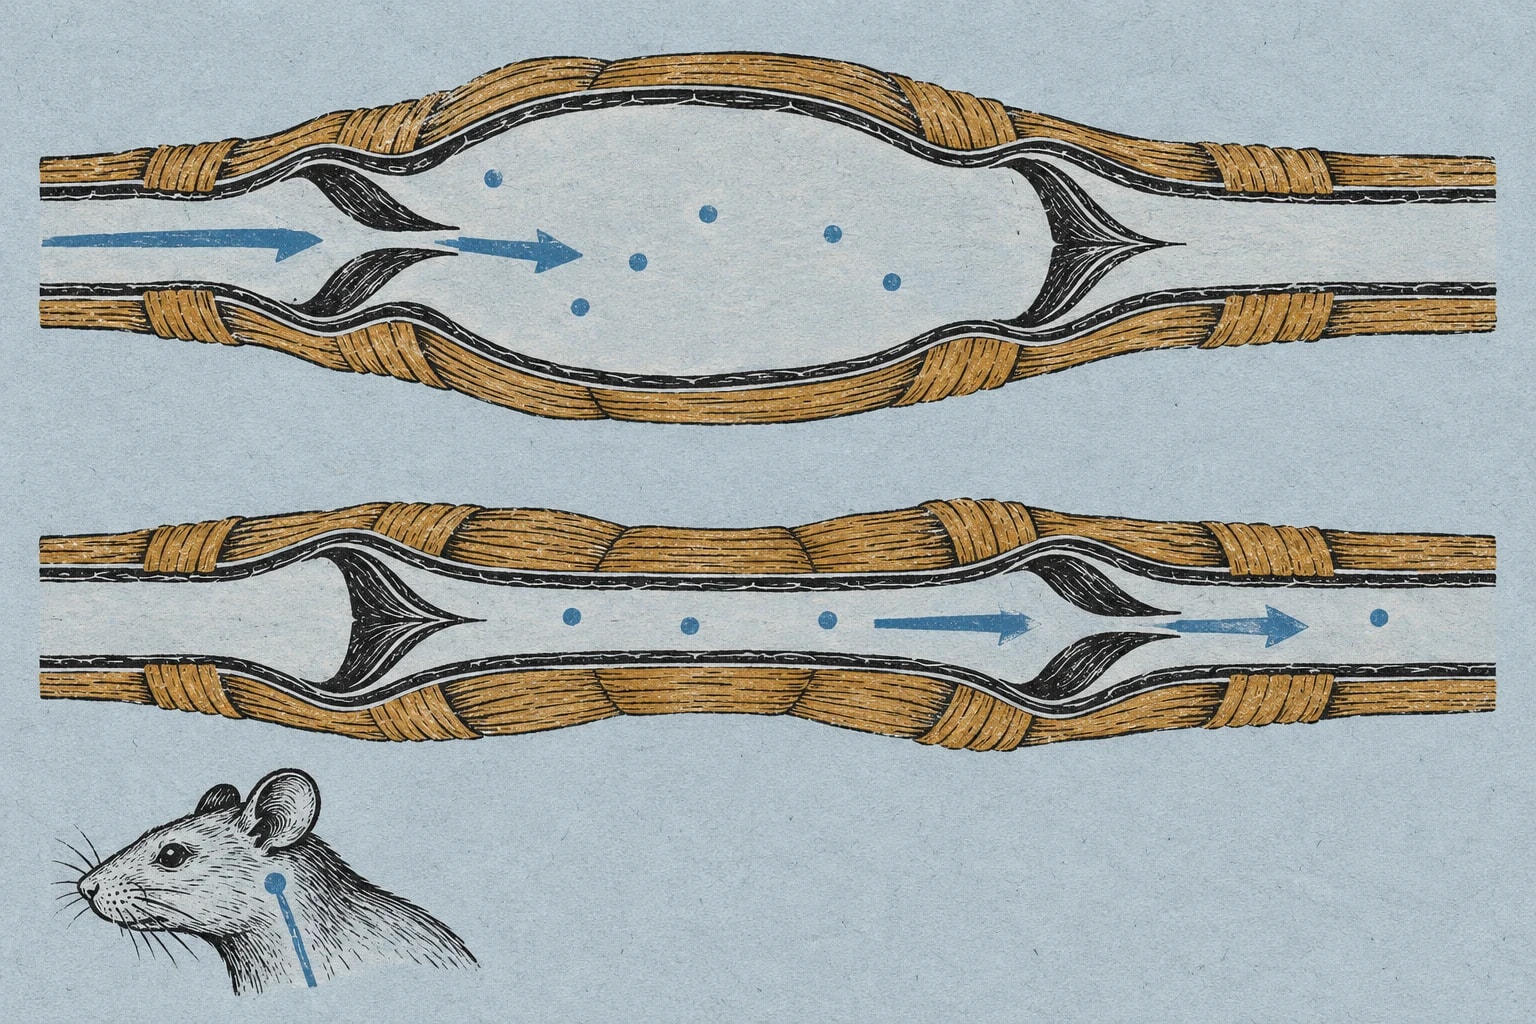
What this could change
Help the brain through the vessels of the neck
Cervical vessels offer a site for restoring drainage from the brain through an intervention outside brain tissue. If improved drainage also protects brain function, this could become a distinct approach to treating age-related impairment. So far, restoration of fluid transport has been demonstrated in aged mice; benefit to brain function remains to be established.
What is known
Local treatment improved cervical lymphatic pumping and cerebrospinal-fluid drainage in old mice.
What to test
Does better drainage also improve brain function and the benefit of intracellular recycling? Faster tracer passage alone establishes improved transport.
What the studies establish
Du and colleagues directly imaged fluid movement in mouse cervical lymphatic vessels. Their pumping weakened with age. Applying prostaglandin F2α directly to the vessels increased contractions and improved drainage of tracers from cerebrospinal fluid in old mice. This demonstrates a reversible transport deficit at a site outside brain tissue.
- Du et al., Nature Aging, 2024Full text available
A separate study by Da Mesquita and colleagues enhanced lymphatic vessels in the brain's membranes using the growth factor VEGF-C. Old mice improved in spatial-learning tasks; tying off the downstream drainage route abolished the behavioral benefit. This supports testing a link between drainage and brain function, but does not establish the same benefit for the cervical-vessel treatment.
- Da Mesquita et al., Nature, 2018Full text available
What remains to be established
The two studies act on different vessel compartments using different interventions. The proposed combination with intracellular recycling, its durability and its effect on lifespan remain untested here.
The test of scale
From a promising result to a rejuvenation program
A particularly convincing result is improvement that persists after treatment ends. It must preserve several functions, resilience to new stress and, in a separate direct test, healthy lifespan.
More on the evaluation criteriaCandidates set aside
These candidates remain in the research record. The reasons below describe their fit for this collection, rather than a verdict that the underlying idea is false. A revised experiment or new evidence can justify reconsideration.
View selection decisions
Can replacing biased blood-forming stem cells alone deliver lasting benefits against aging?
The proposed benefit does not meet the collection’s criteria for breadth and causal focus.
Decision recorded 2026-10-02 05:25 UTC
Local heat buildup may raise oxygen demand and cause hypoxia in the aging kidney
The proposed benefit does not meet the collection’s criteria for breadth and causal focus.
Decision recorded 2026-10-01 04:20 UTC
Delayed immune clearance may drive recurring waves of senescent cells and tissue damage
The proposed benefit does not meet the collection’s criteria for breadth and causal focus.
Decision recorded 2026-10-01 02:45 UTC
Destroying latent viral genomes may reduce recurring damage across organ systems
The proposed benefit does not meet the collection’s criteria for breadth and causal focus.
Decision recorded 2026-10-01 01:00 UTC